
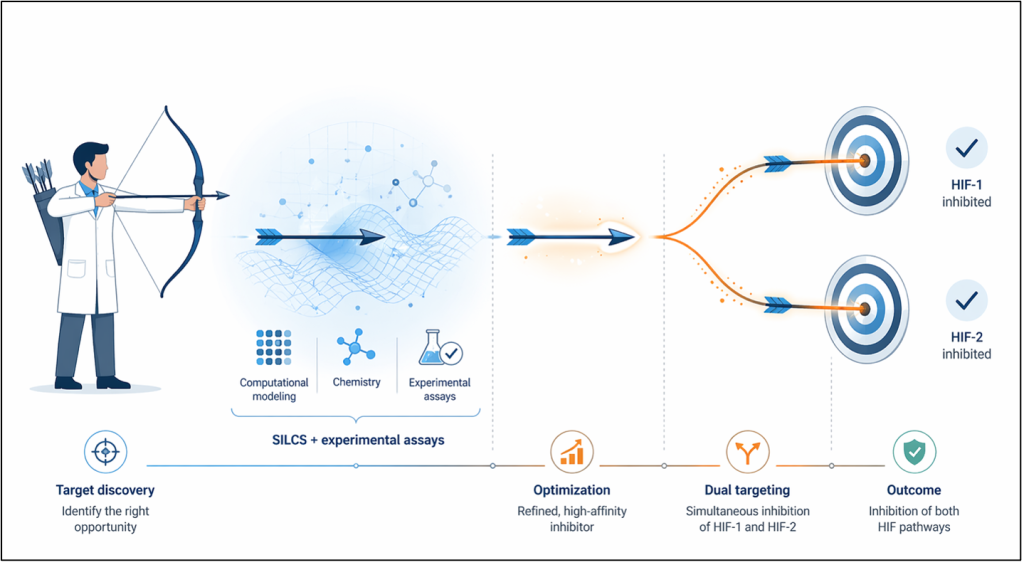
Targeting transcription factors that act as master regulators of cancer progression has long been an important goal in oncology drug discovery. Among the most prominent of these are hypoxia-inducible factors 1 and 2 (HIF‑1 and HIF‑2), which orchestrate broad transcriptional programs that drive tumor growth, angiogenesis, metastasis, and immune evasion. Despite their importance, effectively targeting HIFs, particularly both isoforms simultaneously, has remained a formidable challenge.
Researchers from Johns Hopkins University and the University of Maryland School of Pharmacy recently made strides toward tackling that challenge.
In a study published in the Journal of Experimental Medicine, they reported the discovery of a new class of dual HIF‑1/2 inhibitors that show striking efficacy in multiple cancer models, especially when combined with immune checkpoint blockade. A central component of this work was the application of Site‑Identification by Ligand Competitive Saturation (SILCS), which enabled a focused, probability-driven approach to compound selection and helped bridge computational discovery with robust experimental validation.
Addressing an Elusive Target Class
HIF‑1 and HIF‑2 are transcription factors activated under hypoxic conditions, a hallmark of rapidly growing solid tumors. Their activity regulates hundreds of downstream genes involved in metabolic adaptation, vascularization, and suppression of antitumor immunity. While selective HIF‑2 inhibition has achieved clinical success, notably with belzutifan, accumulating evidence suggests simultaneous inhibition of HIF‑1 and HIF‑2 may be required to more effectively counter tumor progression and resistance mechanisms.
However, transcription factors have historically been viewed as difficult targets for small-molecule therapeutics, in part due to the absence of well-defined binding pockets and the vast chemical space traditionally required to identify viable ligands.
A SILCS-Guided Computational Strategy
To overcome these challenges, Alexander MacKerell Jr., PhD, and colleagues in the Computer-Aided Drug Design Center at the University of Maryland Baltimore School of Pharmacy applied SILCS, a structure-based computational methodology designed to map functional group binding affinities across the full protein surface.
Rather than relying on exhaustive high-throughput screening, the SILCS approach enabled identification of regions on HIF‑2 with a high probability of small-molecule binding. This approach, dramatically accelerated the discovery process while increasing the likelihood of success.
“The SILCS approach enabled the selection of compounds with a high probability of binding to HIF‑2, allowing experimental efforts to focus on testing hundreds, rather than millions, of chemical compounds,” said Alex MacKerell.
Notably, while the initial computational studies focused on HIF‑2, subsequent experimental work revealed compounds capable of engaging both HIF‑1 and HIF‑2, triggering their degradation and suppressing downstream oncogenic signaling.
From Computational Predictions to Biological Validation
Building on the SILCS-guided compound selection, collaborators at Johns Hopkins University led by Gregg L. Semenza, MD, PhD, performed extensive biological characterization. The identified compounds demonstrated potent inhibition of HIF signaling across a range of cancer cell lines and reduced angiogenesis and invasiveness in vivo.
In mouse models of breast, colorectal, melanoma, and prostate cancer, the dual HIF inhibitors alone suppressed tumor growth. The most striking results emerged when the compounds were combined with immune checkpoint inhibitors, such as anti‑CTLA‑4 or anti‑PD‑1. In more than half of treated animals, including in models resistant to immunotherapy alone, tumors were eliminated and remained absent upon rechallenge.
Mechanistic studies showed that the combination therapy reshaped the tumor microenvironment by reducing immunosuppressive cell populations and increasing cytotoxic T cells and natural killer cells, offering a clear biological rationale for the observed synergy.
Implications for Drug Discovery
This work provides a compelling demonstration of how structure-based computational methods can expand the range of tractable targets and enable discovery strategies that would be impractical using traditional screening approaches alone. By guiding compound selection toward high-probability binding interactions early in the pipeline, SILCS played a key role in translating computational insight into therapeutically meaningful outcomes.
Beyond the specific context of HIF biology, the study highlights a broader principle: rational, physics-based mapping of protein–ligand interactions can enable efficient exploration of challenging target space and facilitate productive integration with experimental and translational research.
Read the Full Study
The full findings are detailed in the April 2 publication in the Journal of Experimental Medicine:
Targeting conserved domains of hypoxia-inducible factors reveals potent dual HIF‑1/2 inhibitors with broad antitumor activity
Learn More about SILCS
If you are interested in exploring how the SILCS platform can support your drug discovery program, please contact us at info@silcsbio.com

